Prediction of single cell gene expression

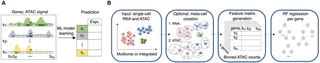
Learning the relationship between gene-regulatory regions from epigenome and transcriptome data measured for single-cell is hard. We have developed an automated Nextflow pipeline that addresses the noise and sparsity of single-cell data by aggregating individual cells into metacells. We show that analysis on metacell level often leads to superior models and we illustrate the use of these models on different datasets.
Linked publications
(2026).
Predicting gene-specific regulation with transcriptomic and epigenetic single-cell data.
Bioinformatics.

Authors
Laura Rumpf
(she/her)
PhD student
I’m a computer scientist by training who found my way into bioinformatics during my master’s. My work focuses on understanding enhancer–gene regulation using machine learning and single-cell transcriptomic and epigenomic data. I’m particularly interested in how computational models can help uncover regulatory mechanisms and translate these insights into disease-related questions in computational medicine. Outside the lab, I’m usually fueled by coffee and a slightly too serious interest in latte art.